 Get Insights

How to Plan Medical Equipment Validation and Qualification for ISO 13485 Readiness
How to Plan Medical Equipment Validation and Qualification for ISO 13485 Readiness
ISO 13485 readiness depends on more than a controlled procedure and a folder of completed forms. Auditors, notified bodies, customers, and internal quality leaders need objective evidence that equipment performs safely, consistently, and for its intended use. That evidence has to connect back to the product, the process, the patient or user risk, and the decisions made when something falls outside acceptance limits.
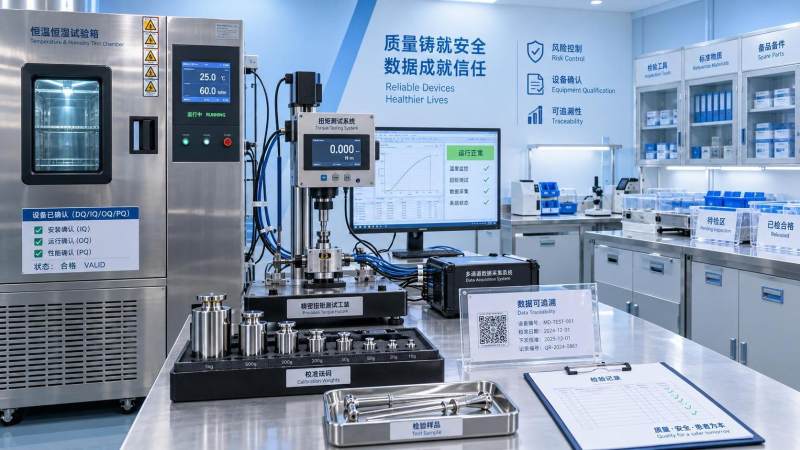
This is where medical equipment validation qualification support becomes practical rather than administrative. A strong program turns broad requirements into a defensible plan: which equipment needs qualification, what conditions must be challenged, who can approve deviations, how records remain traceable, and when requalification is triggered. The detail matters. A temperature-controlled chamber used for stability work, a torque-controlled assembly station, and an inspection camera may all be “equipment,” but they do not carry the same risk or require the same evidence.
One frequent mistake is treating every asset as if it needs the same IQ/OQ/PQ package. The opposite mistake is relying on a supplier’s brochure, factory acceptance test, or calibration certificate as proof that the equipment works in the manufacturer’s actual environment. Neither approach stands up well when a quality issue emerges. The useful question is simpler: what must this equipment reliably do, under what conditions, and what could happen if it does not?
Start by separating equipment qualification from device and process validation
The terminology is often blurred, especially in growing MedTech organizations. Qualification generally establishes that a facility, utility, system, or item of equipment is installed and operating as required. Validation establishes documented evidence that a process, software application, or product-related activity consistently achieves its intended result. In practice, the two overlap: an assembly process cannot be convincingly validated if the dispensing machine, curing oven, vision system, or test fixture supporting it is not fit for use.
For ISO 13485:2016 readiness, the boundary is important. Clause 7.5.6 addresses validation of processes for production and service provision when output cannot be fully verified by subsequent monitoring or measurement. Clause 7.6 addresses control of monitoring and measuring equipment. Design validation has a different role within design and development controls. A qualification plan should show how these areas interact without pretending they are interchangeable.
For example, a tensile tester may require calibration and suitability checks because its readings support product acceptance. A heat-sealing process may need process validation because seal integrity cannot always be assured by inspecting every finished pouch. The sealing machine itself may need installation and operational qualification as part of the evidence base. The equipment file should support the process file; it should not become a disconnected documentation exercise.
Build the scope around patient risk and product decisions
Before writing protocols, create an equipment inventory that is meaningful to quality. Many organizations already have an asset register, but it often mixes office hardware, maintenance tools, production equipment, laboratory instruments, and software-enabled test systems without showing their quality impact. The qualification scope should identify the equipment’s intended use, the process or test it supports, the controlled parameters, associated software or utilities, and the product decisions influenced by its output.
Then apply a documented risk assessment. The method can fit the organization’s quality system, but it should consider more than the chance of equipment failure. Consider whether a failure could escape detection, whether it affects sterile barrier integrity, dosage, electrical safety, dimensional conformance, data integrity, or traceability, and whether an independent check exists downstream. An analytical balance used to prepare a critical reagent deserves a different level of scrutiny from a label printer used only for internal storage locations.
This risk-based distinction makes planning more efficient. It also helps when auditors ask why one asset has an extensive operational challenge study while another has a documented commissioning review, calibration status, and limited functional verification. The answer should be found in the intended use and risk record—not in a statement that the team “considered it low risk.”
Define intended use before selecting test cases
A user requirements specification, often called a URS, is not a purchasing wish list. It is the anchor for qualification. It should say what the equipment must accomplish in the organization’s own operating context. “The system shall be accurate” is too vague to test. “The test system shall prevent unauthorized alteration of approved test methods” is clearer, provided the requirement is relevant to the intended use and supported by a defined verification approach.
Requirements should cover the failure points that matter in real use: operating range, resolution or repeatability where relevant, environmental conditions, alarm behavior, power interruption recovery, data storage, user permissions, interfaces, cleaning constraints, and maintenance access. For a device connected to manufacturing execution software or a laboratory information system, the interface is often more fragile than the instrument itself. A qualification plan that tests the machine in isolation but not the transfer, storage, and review of its data can leave a major gap.
Supplier documentation can be useful here, but it should not dictate the requirements. Marketing claims and standard factory tests may demonstrate capability under controlled conditions. They do not automatically demonstrate that the same capability is maintained after installation, configuration, network connection, local utility variation, or routine operator use.
Use protocols that test the edges, not just the happy path
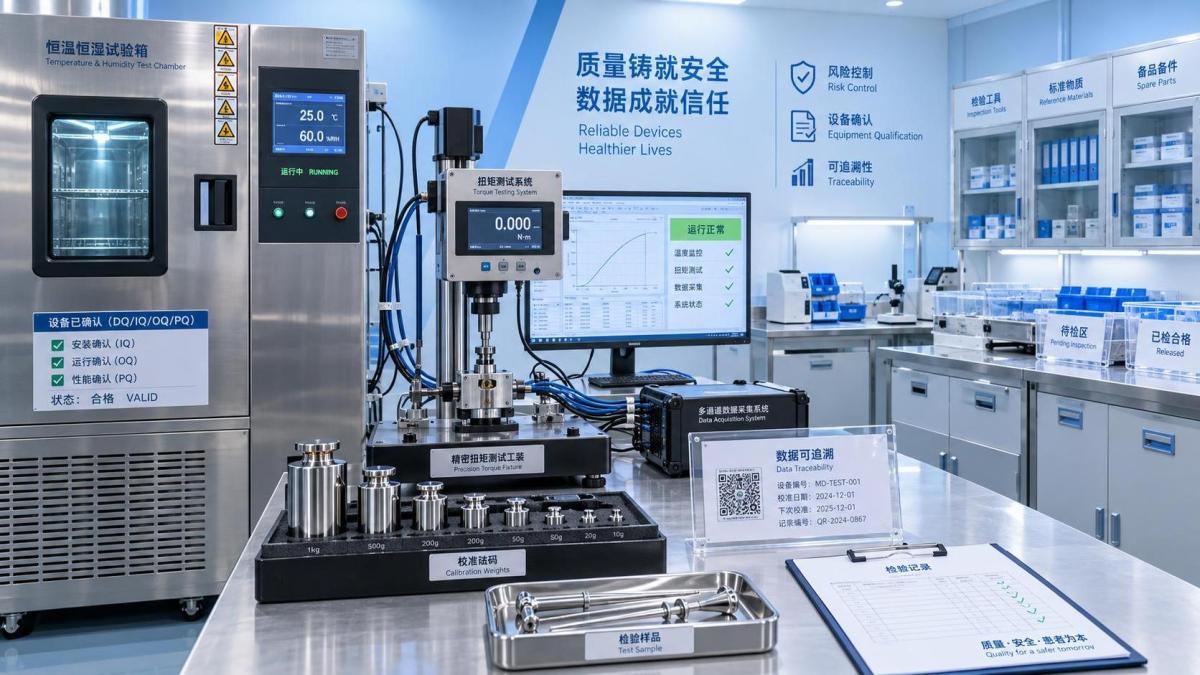
Qualification protocols should be executable by a competent person who was not involved in writing every line. That is a good practical test of clarity. Each test needs a stated purpose, preconditions, step-by-step actions, expected results, actual results, objective evidence, and acceptance decision. Protocols should identify the approved configuration, instrument IDs, software versions, reference standards, and any required environmental conditions.
Installation qualification commonly confirms that the delivered system matches the approved configuration and is installed correctly: utilities, connections, components, manuals, safety features, software versions, calibration status, and preventive maintenance requirements. It sounds straightforward, but configuration drift often begins here. If a supplier substitutes a component, installs an unapproved firmware version, or leaves a service account active, those details should not be discovered months later during an investigation.
Operational qualification should challenge the defined operating conditions and foreseeable abnormal conditions. Do not limit the exercise to a single nominal setpoint if the process uses a range. Test alarms, interlocks, parameter limits, recovery behavior, failed sensor conditions where safely feasible, and user-role restrictions. The aim is not to force equipment to fail theatrically; it is to show that controls work when routine conditions are less than perfect.
Performance qualification is often the point at which teams become vague. It should demonstrate sustained performance in the intended production or laboratory setting, using representative materials, operators, and operating conditions. The exact approach depends on process risk and the evidence already available. It must also align with the broader validation strategy. A qualified oven does not by itself prove that a bonded medical device meets its cure profile; material behavior, load pattern, fixture design, and downstream testing may all matter.
Treat software, data, and suppliers as part of the equipment system
Modern equipment rarely operates as a standalone mechanical asset. A vision inspection station may classify defects through configured logic. A wearable-sensor test bench may capture signal data through a proprietary application. A laboratory analyzer may calculate results, export files, and maintain audit trails. If the software affects quality records or product acceptance, it needs assessment and validation commensurate with risk under the organization’s software validation procedures.
A common weak point is assuming that vendor qualification covers the full system. Supplier assessment is necessary, especially for specialized test equipment and digital platforms, but a supplier’s quality documentation does not replace site-specific evidence. Review what is available: manuals, calibration certificates, release notes, service reports, known limitations, cybersecurity information where relevant, and change notifications. Then identify what must be independently verified at the user site.
Independent technical benchmarking can be valuable when performance claims are difficult to interpret. In complex MedTech supply chains, organizations such as VitalSync Metrics examine engineering characteristics that promotional material may not explain clearly—from wearable sensor signal quality to material fatigue behavior. That kind of external evidence can inform procurement and risk assessment. It does not replace a manufacturer’s own qualification work, because intended use, configuration, and local process conditions remain specific to the organization.
Make deviations useful rather than embarrassing
A qualification package with no deviations is not automatically a strong package. It may simply mean that the protocol avoided meaningful challenges or that observations were handled informally. Deviations should be recorded when a test does not execute as approved, an expected result is not met, required evidence is missing, or the equipment configuration changes during execution. The record should describe what happened, assess product and quality-system impact, identify the root cause where appropriate, and define the disposition.
Be cautious about closing a deviation with “retest passed.” A successful retest may show that the immediate issue is no longer visible, but it may not explain whether the original result affected prior data, other equipment, released product, or the validity of the protocol. Quality and safety personnel should insist on a proportionate impact assessment. This is particularly important for test systems that have already generated release, in-process, or stability data.
Plan the lifecycle before approval
Qualification is not complete when the final report is signed. The report should summarize executed testing, deviations, unresolved limitations, conclusions, and the approved status of the equipment. More importantly, it should state the controls needed to keep that status meaningful: calibration, preventive maintenance, periodic review, operator training, backup and restore procedures for electronic data, and change control.
Requalification triggers should be defined in advance. Typical triggers include relocation, replacement of critical components, firmware or software updates, changes to utilities, a major repair, revised operating ranges, repeated maintenance failures, or a change in intended use. Not every change requires repeating every qualification test. But the decision should be risk-based, documented, and made before the changed equipment resumes quality-impacting work.
The most audit-ready programs are usually not the largest ones. They are the programs where the asset list, risk assessment, URS, protocols, raw evidence, deviation records, calibration controls, and change history tell one consistent story. If a quality reviewer can follow that story from a product requirement to an equipment setting—and from a failed alarm test to the corrective decision—the organization has built evidence that is useful well beyond an ISO 13485 audit.
Recommended News
- 2026.10.14Clinic Supplier Comparison: Evaluating Cost, Delivery Reliability, and SupportDr. Alistair Thorne (Senior Biomedical Engineer)
- 2026.10.13Engineering Performance Testing Services: Evaluating Test Methods and AccuracyDr. Alistair Thorne (Senior Biomedical Engineer)
- 2026.10.12Preparing for an ISO 13485 Audit: When to Use External QMS SupportDr. Julian Rossi (RehabTech Specialist)
The VitalSync Intelligence Brief
Receive daily deep-dives into MedTech innovations and regulatory shifts.