 Get Insights

Technical Integrity Compliance: How to Document Design Changes
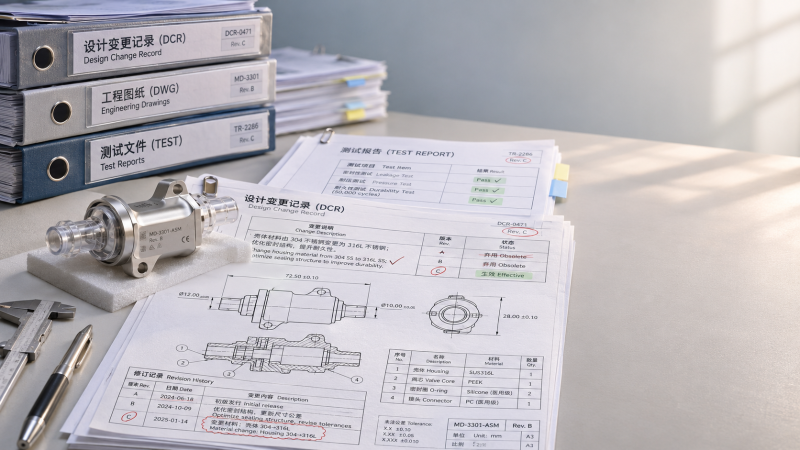
Design changes can improve medical devices, but an incomplete record can conceal new safety risks, break traceability and complicate regulatory review. For quality control and safety managers, technical integrity compliance depends on more than a signed change form: the evidence must show what changed, why it changed, how its effects were assessed and whether the implemented product still meets its requirements.
The practical challenge is connecting a decision made in one place—a supplier notice, software ticket or engineering deviation—to every affected requirement, risk control, test and released configuration. The following approach helps build that connection while distinguishing good documentation practice from obligations that depend on the product and applicable regulations.
Start with the regulatory and technical baseline
Before evaluating a proposed change, identify the approved baseline: the device configuration, specifications, software version, manufacturing process and documents currently authorized for use. Without it, reviewers cannot reliably determine the scope of the change or reproduce the evidence supporting the previous design.
For medical devices placed on the EU market, Regulation (EU) 2017/745 (MDR) requires manufacturers to maintain a quality management system that addresses, among other matters, design and development, risk management and product realization. Regulation (EU) 2017/746 (IVDR) contains corresponding obligations for in vitro diagnostic devices. Both regulations also require technical documentation to be kept up to date. See MDR Article 10 and Annex II, and IVDR Article 10 and Annex II: https://eur-lex.europa.eu/eli/reg/2017/745/oj and https://eur-lex.europa.eu/eli/reg/2017/746/oj.
ISO 13485:2016 addresses design and development changes within a medical-device quality management system; ISO 14971:2019 addresses lifecycle risk management. Obtain the applicable standards and assess them against the organization’s procedures and product scope. A standard is not a substitute for the law applicable in each market. Sources: https://www.iso.org/standard/59752.html and https://www.iso.org/standard/72704.html.
Build a change record that can stand on its own
A reviewer should be able to reconstruct the decision without relying on an engineer’s memory or an email thread. Use a unique change identifier and link the record to controlled evidence rather than copying uncontrolled attachments into multiple systems.
- Define the change. Identify the affected product, variants, components, processes and document revisions. State the current and proposed configurations, including software or firmware versions where relevant.
- Record the reason. Link to the initiating event, such as a nonconformance, complaint investigation, supplier notification, corrective action or design improvement. Separate verified facts from assumptions.
- Establish the boundaries. Specify what is not changing as well as what is. This helps reviewers challenge an impact assessment that overlooks an interface or dependent component.
- Document the assessment. Identify affected requirements, hazards, risk controls, tests, labeling, manufacturing instructions and regulatory submissions. Record why an area is unaffected when that conclusion matters.
- Preserve the decision. Identify reviewers, their functions, approval dates, conditions and any rejected alternatives. Retain links to the evidence used at approval.
- Control implementation. Record the effective date or other implementation trigger, affected inventory, training and final released revisions. Close the record only when required actions and evidence are complete.
These are practical record-design elements, not a claim that every field is expressly prescribed in the same form by every regulation. Adapt the detail to the change and the applicable quality system.
Assess impact before deciding how much testing is enough
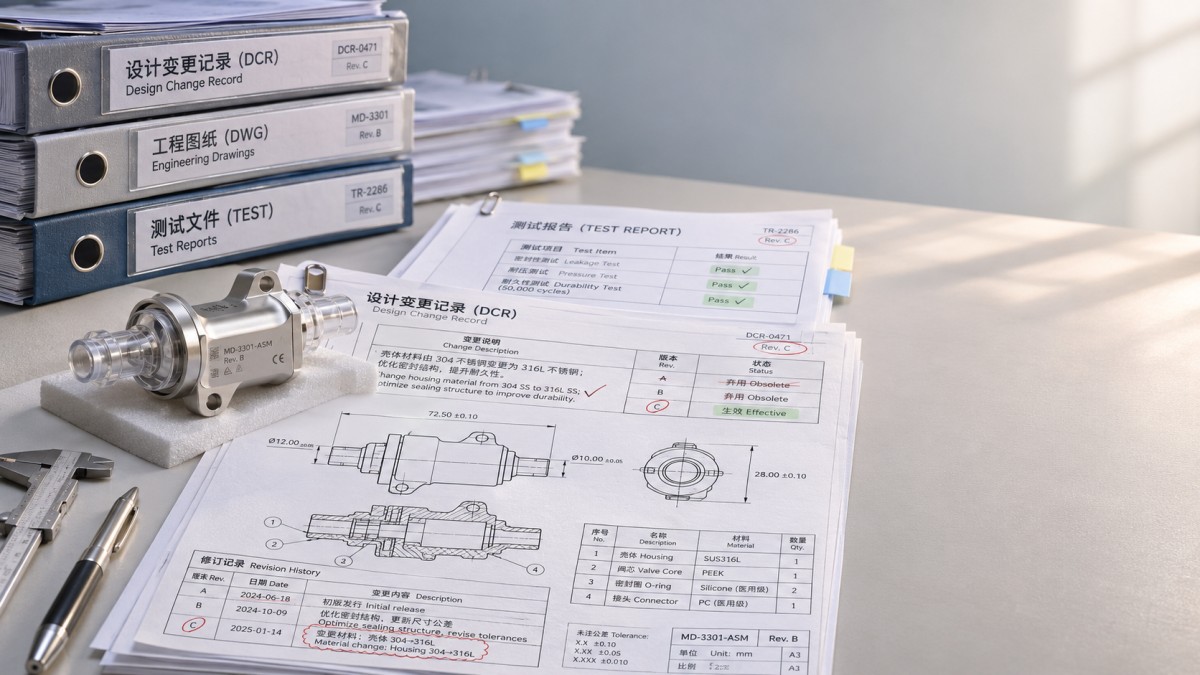
A seemingly minor substitution can change a product’s performance or its manufacturing controls. For example, a revised polymer grade may retain the same nominal dimensions while affecting a material specification, a critical process setting or the evidence supporting a finished-device requirement. The record should show the reasoning, not merely label the change “minor.”
Engage the functions that own the affected evidence. Engineering may understand the interface, while quality, manufacturing, clinical or performance specialists, and regulatory affairs may identify consequences outside the drawing. For an IVD, consider whether a change affects specimens, reagents, instruments, software or the performance claims supported by the existing evidence.
Translate risk into a verification and validation plan
Verification asks whether design outputs meet specified inputs; validation considers whether the resulting design meets user needs and intended uses. The change record should identify which activities are needed, their acceptance criteria, the responsible owner and the approved protocol before results are judged. These concepts are reflected in FDA design-control requirements and guidance: 21 CFR 820.30 and FDA’s Design Control Guidance, available at https://www.ecfr.gov/current/title-21/chapter-I/subchapter-H/part-820 and https://www.fda.gov/regulatory-information/search-fda-guidance-documents/design-control-guidance-medical-device-manufacturers.
Do not assume that rerunning one historical test proves the entire change is acceptable. Explain why selected tests address the affected requirements and risks, why a sample or configuration is representative, and whether existing evidence remains applicable. If a test is not repeated, document the technical justification and its limitations. Investigate failures and deviations; do not close them by replacing an unfavorable result with a later passing result alone.
FDA’s Quality Management System Regulation took effect on February 2, 2026, incorporating ISO 13485:2016 by reference with additional requirements. Check the current regulation and FDA guidance when applying US requirements rather than relying on an older procedure’s section numbering. Source: https://www.fda.gov/medical-devices/quality-and-compliance-medical-devices/quality-management-system-regulation-qmsr.
Maintain traceability from decision to released configuration
A traceability matrix need not be a separate spreadsheet if the quality system can produce the same clear relationships. What matters is that a reviewer can move in both directions: from the change to the affected requirement and test, and from a released product version back to the evidence that supports it.
- Link the proposal to the baseline. Cite controlled document identifiers and revisions, not only filenames or shared-drive locations.
- Link impacts to requirements and risks. For each affected item, identify the disposition: updated, unchanged with justification, or subject to further assessment.
- Link tests to acceptance decisions. Reference approved protocols, reports, deviations and the configuration actually tested.
- Link approval to release. Identify the authorized approvers and the production, software or document configuration that may be implemented.
- Confirm implementation. Reconcile the approved change with the released bill of materials, manufacturing instructions, inspection controls and applicable labeling.
Pay particular attention to timing. A design may be approved before a supplier has delivered the revised component, while old and new stock may coexist. Define how affected lots, serial numbers or other identifiers will be controlled where applicable, and what happens to work in progress. An effective-date field without an implementation plan does not demonstrate which configuration was actually supplied.
Decide whether external notification or certification action is needed
Do not equate an internal change classification with a regulatory conclusion. Under the MDR and IVDR, relevant notified body obligations depend on the conformity assessment route and certificate conditions. The regulations include provisions requiring manufacturers to inform the notified body of planned substantial changes to an approved quality management system or covered device range under the applicable assessment procedures; other route-specific provisions may also apply. Review the relevant annex and the agreement with the notified body before implementation. Sources: MDR Annex IX and IVDR Annex IX at the EUR-Lex links above.
EU transitional devices require particular care: whether a change is “significant” for transitional provisions is a specific regulatory assessment, not simply the organization’s internal risk rating. Consult the applicable current MDCG guidance, including MDCG 2020-3 for MDR legacy devices and MDCG 2022-6 for IVDR legacy devices, and retain the version used in the decision. Source: European Commission medical device guidance, https://health.ec.europa.eu/medical-devices-sector/new-regulations/guidance-mdcg-endorsed-documents-and-other-guidance_en.
Record the markets reviewed, the applicable route, the rationale, the decision owner and any required communication. If notified body input is needed, preserve the correspondence and distinguish a submitted notification from an authorization or approval actually received. A regulatory assessment should be revisited if the technical scope changes during development.
Review the evidence before closing the change
Quality review should test the integrity of the record, not just the presence of signatures. Ask whether the stated change matches the released configuration, whether open test deviations were resolved, and whether risk controls remain traceable to objective evidence. Check that affected procedures, supplier controls and training are complete, or that any permitted outstanding actions have explicit owners and due dates.
Finally, define how post-implementation information will feed back into the assessment. Complaints, nonconformities, service findings and supplier issues may reveal effects that were not apparent during verification. Where the change alters risk conclusions or technical documentation, update those controlled records rather than leaving the change order as the only source of truth.
Frequently asked questions
Does every design change require a complete retest?
Not necessarily. The organization should determine the verification and validation needed for the affected requirements and risks, then document its rationale. Existing evidence may remain useful if its applicability to the new configuration is justified; a blanket exemption based only on the word “minor” is not a sufficient technical explanation.
Can a supplier’s change notice serve as the design change record?
It can be supporting evidence, but it normally does not replace the manufacturer’s assessment. The manufacturer needs to determine how the supplier change affects its own device, requirements, risks, manufacturing controls and regulatory obligations.
Who should approve a change?
Approvals should follow the organization’s controlled procedure and involve the functions responsible for the affected decisions. At a minimum, reviewers need the authority and competence to assess the relevant technical, quality and regulatory consequences; the exact roles depend on the change and the applicable quality system.
Recommended News
- 2026.11.22MedTech Benchmarking: Which Metrics Reveal Operational Performance?Dr. Alistair Thorne (Senior Biomedical Engineer)
- 2026.11.21Study Finds Declining Device Failure Transparency, Raising Questions About FDA OversightDr. Julian Rossi (RehabTech Specialist)
- 2026.11.21Technical Integrity Compliance: How to Document Design ChangesDr. Julian Rossi (RehabTech Specialist)
The VitalSync Intelligence Brief
Receive daily deep-dives into MedTech innovations and regulatory shifts.